Introduction to Molecular Simulations
Molecular Simulation is the science of simulating the motions and interactions of atoms and molecules to understand microscopic behavior.
For engineers and materials scientists, it acts as a “Computational Microscope.” It allows us to zoom in beyond what is visible in a lab to understand why a material fails, how a drug binds, or what drives a chemical reaction.
Where are they used?
Molecular simulations have moved from theoretical physics into standard engineering workflows.
Mechanical Engineering: Understanding fracture mechanics at the crack tip, friction (tribology), and heat transfer in nanomaterials.
Chemical Engineering: Predicting transport properties (diffusion, viscosity) for new fluids, designing catalysts, and studying interfacial phenomena.
Materials Science: Designing battery electrolytes, predicting crystal structures, and developing polymers with specific elasticity.
Biomedical Engineering: Protein folding dynamics, drug discovery (docking), and designing lipid nanoparticles for drug delivery.
Why are they becoming important?
Two major trends are converging to make this a golden age for molecular simulation:
The Breakdown of Empirical Laws: As we engineer devices at the nanoscale (micro-fluidics, NEMS, advanced drug delivery), continuum theories like Navier-Stokes or continuum mechanics often break down. We need atomistic detail.
The Rise of Compute: Moore’s Law and the advent of GPU computing have allowed us to simulate systems that were impossible 10 years ago.
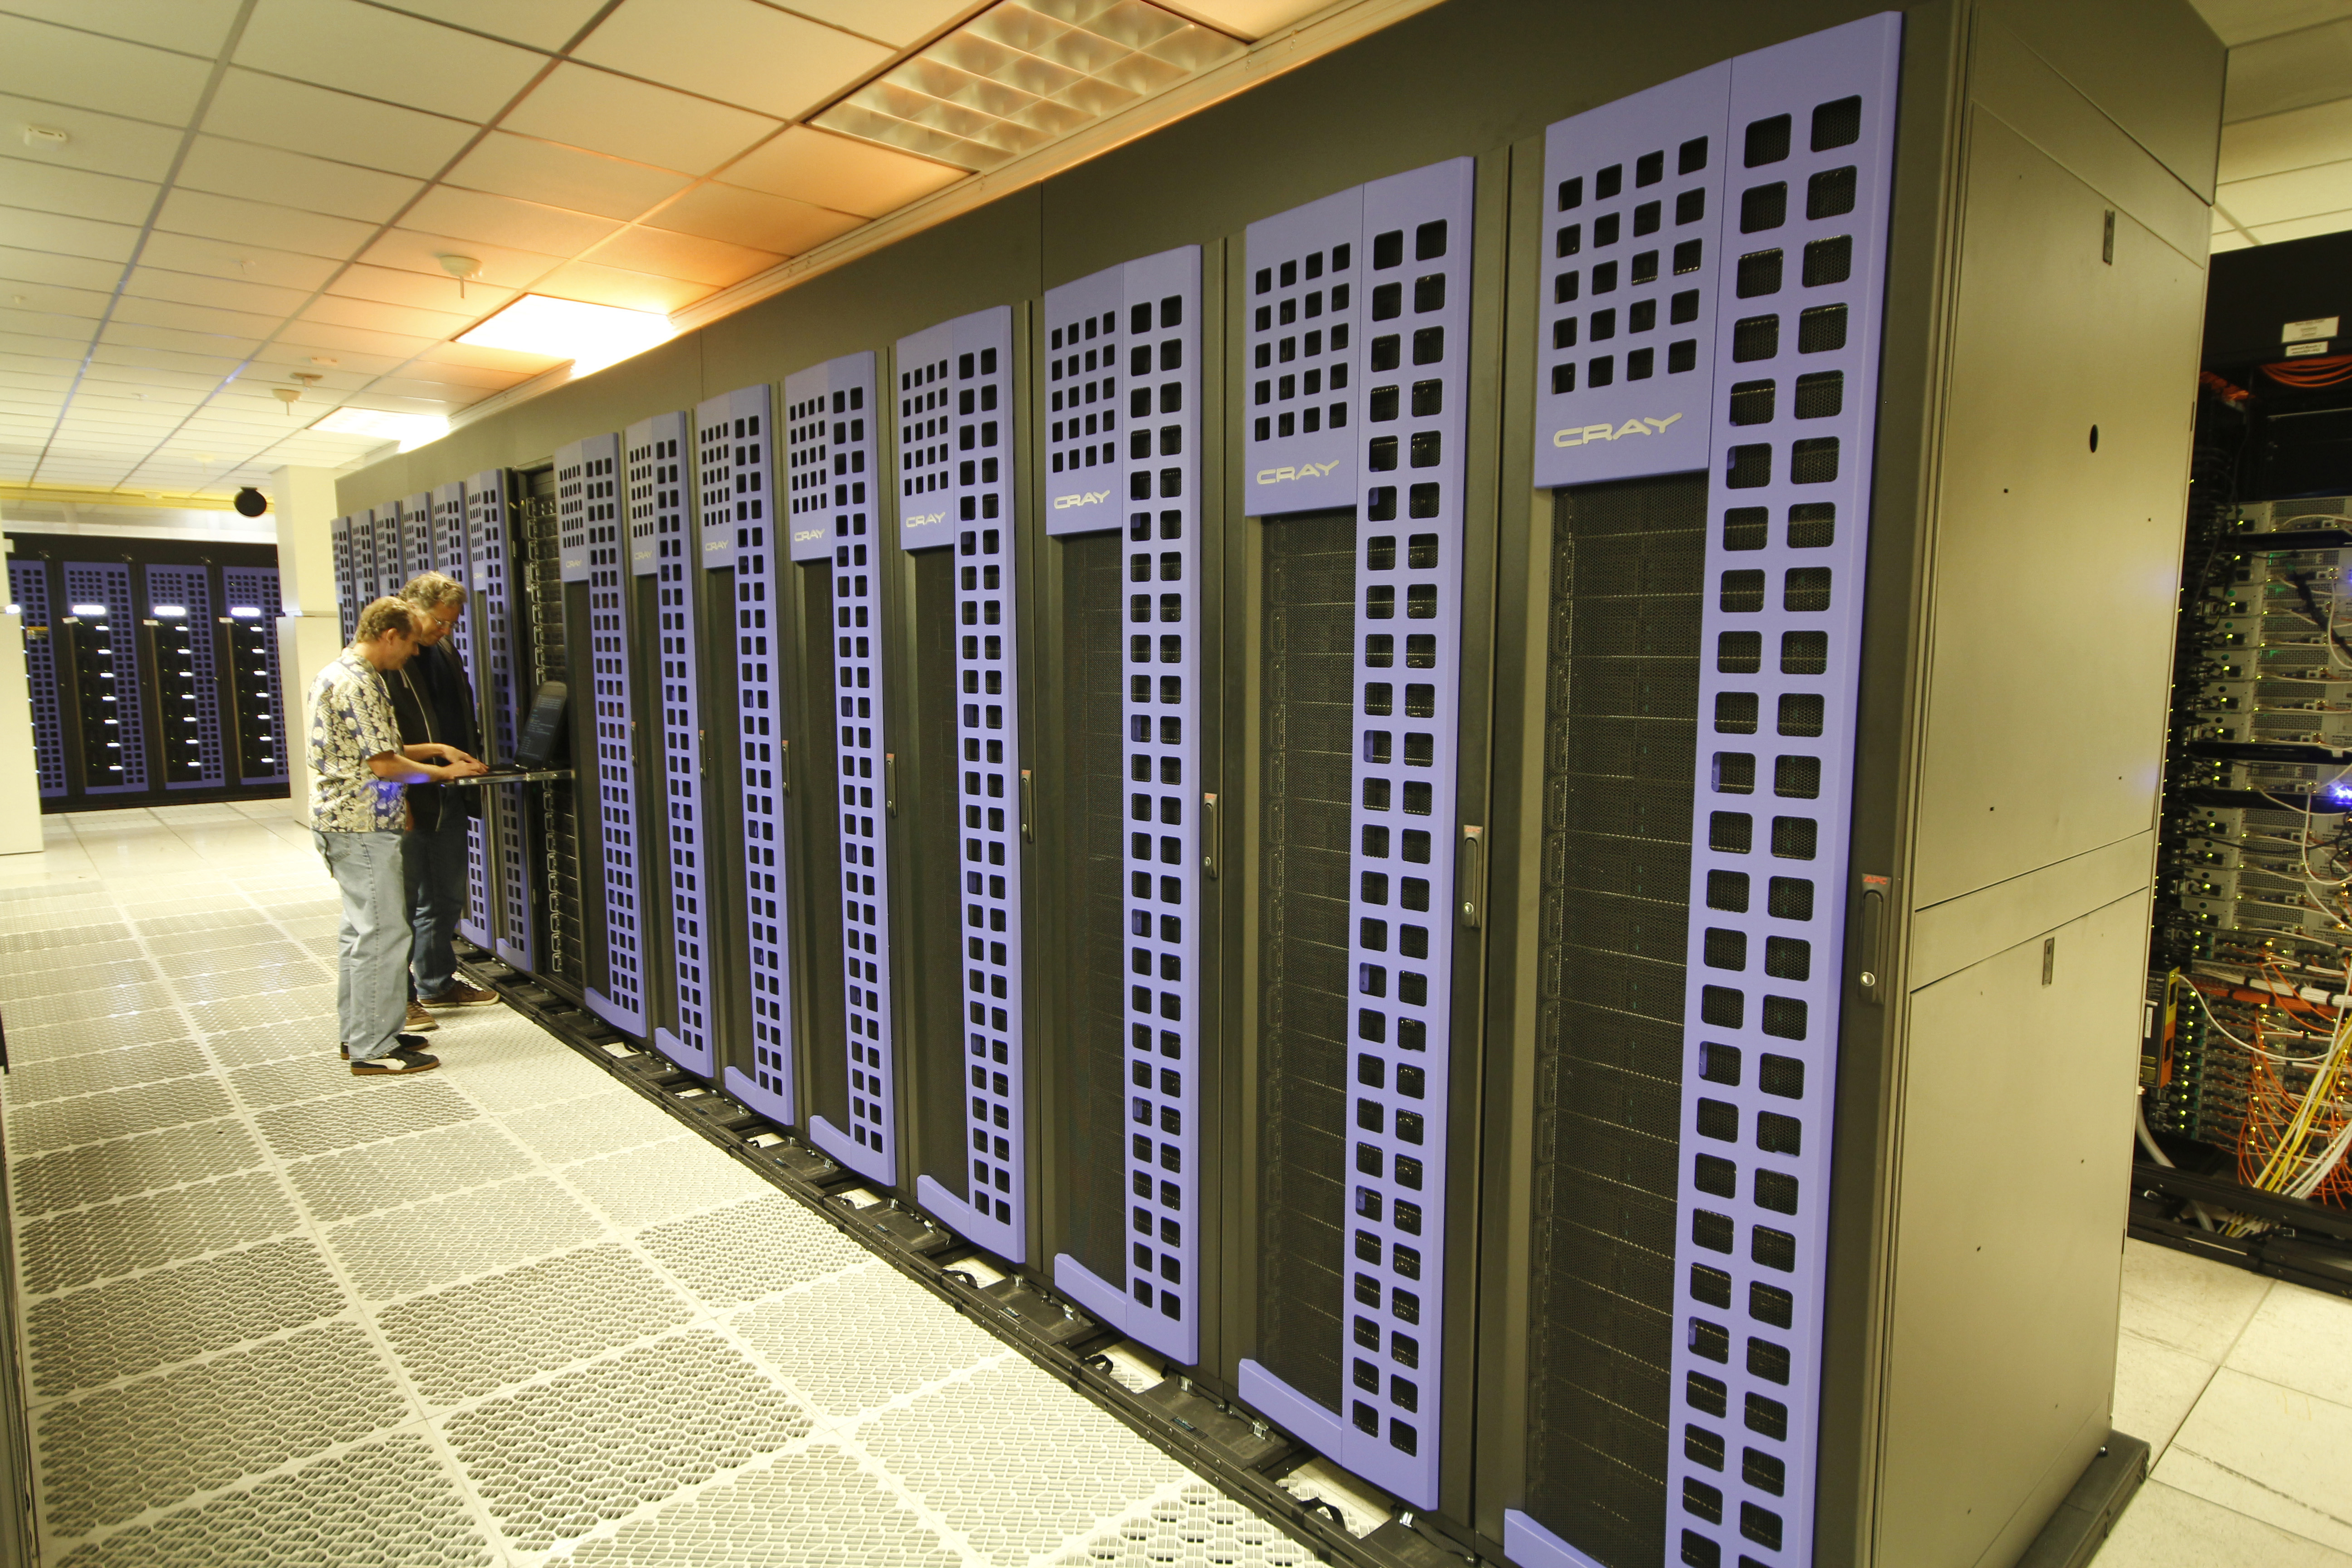
Figure 1:High Performance Computing (HPC) clusters are the “wind tunnels” of molecular engineering.
How are they run? (The Role of HPC)
A typical molecular dynamics simulation involves calculating the forces between every pair of atoms in a system.
The Scale Problem
If you have a system with atoms, a naive calculation of forces requires checking every pair, leading to interactions. Even with clever algorithms (bringing it down to ), this is computationally expensive.
Time Step: To capture atomic vibrations, we move atoms in steps of 1 femtosecond (10-15 s).
Target Time: To see a protein fold or a crack propagate, we need microseconds (10-6 s).
The Math: You need 109 (one billion) steps to simulate one microsecond.
Why we use Supercomputers (HPC)
You cannot run meaningful production simulations on a laptop. We use High Performance Computing (HPC).
Parallelization: We split the simulation box into chunks (Domain Decomposition).
MPI (Message Passing Interface): Different processors “talk” to each other to hand off atoms that move across boundaries.
GPUs: Modern codes (like GROMACS, LAMMPS, AMBER) use Graphics Processing Units to calculate forces 100x faster than standard CPUs.